
Companies that design and produce medical devices operate within technical and regulatory frameworks that shape product development, manufacturing, and market access. Manufacturers translate clinical and user needs into engineered products, then apply process controls, materials selection, and documentation to meet applicable safety and performance expectations. This work typically involves multidisciplinary teams—engineering, quality, regulatory, and production—working to align designs with recognized technical standards, risk analyses, and manufacturing capabilities. The result is a product pathway that emphasizes traceability, repeatability, and documented evidence that relevant requirements were considered during development and production.
Regulatory authorities and consensus standards often prescribe the structure of product files, testing evidence, and quality systems that manufacturers may use to show conformity. Standards such as ISO 13485 for quality management and IEC technical standards for electrical safety are commonly referenced in industry guidance, while national and regional regulators define conformity assessment routes that can vary by product classification and intended use. Manufacturers typically maintain design history files, device master records, and batch-level records, and they may engage third-party testing laboratories to validate performance against published norms and regulatory expectations.
Manufacturers often use risk management frameworks to connect design decisions with testing and manufacturing controls. A risk-based approach may include hazard identification, severity and probability assessments, and determination of risk-control measures. This process typically informs verification activities—demonstrations that design inputs produce intended outputs—and validation activities where intended use in realistic conditions is assessed. Risk files and traceability matrices commonly accompany device dossiers so that reviewers can follow how identified hazards were mitigated through design features, labeling, or instructions for use.
Supply chain controls and supplier qualification are frequently central to production consistency. Manufacturers may assess suppliers for capability, audit critical suppliers, and define incoming inspection protocols for key components. For devices that include off-the-shelf or custom software, source code controls, configuration management, and documented testing protocols can be particularly important. Where sterilization or sterile packaging is required, validation of sterilization cycles and packaging integrity testing are often necessary elements in product release criteria and regulatory submissions.
Regulatory compliance pathways typically require comprehensive technical documentation that links requirements to evidence. Such documentation can include design specifications, test reports, clinical literature summaries if applicable, manufacturing records, and post-market surveillance plans. Conformity assessment routes vary: some devices may follow self-declaration processes, while others require third-party assessment or clinical data. Manufacturers often prepare clear device descriptions and intended-use statements because regulatory classification and the applicable conformity assessment depend on how the device will be used and by whom.
Post-market monitoring and corrective processes are integral to maintaining compliance after product release. Manufacturers commonly implement systems for complaint handling, adverse event reporting, trend analysis, and periodic review of device performance. If a nonconformity or safety concern emerges, established corrective and preventive action (CAPA) procedures may be used to investigate root causes and implement changes to design, manufacturing, or labeling. These post-market inputs can also inform periodic updates to risk assessments and technical documentation, supporting ongoing conformity.
In summary, companies that produce medical devices coordinate design, process control, testing, and documentation to meet technical standards and regulatory expectations. Workflows typically emphasize traceability and risk management, with supply chain oversight and post-market surveillance forming continuing obligations. The next sections examine practical components and considerations in more detail.
Design activities for regulated medical devices often begin with defined user needs and intended use statements that shape design inputs. Manufacturers typically translate these inputs into specifications for performance, materials, and safety. Risk management is commonly integrated into design processes, allowing teams to identify hazards, estimate associated risks, and apply controls or mitigations. Traceability matrices may link each design input to verification and validation activities. Insight from human factors or usability engineering can be used where user interaction affects safety or performance, and this may inform labeling and instructions.
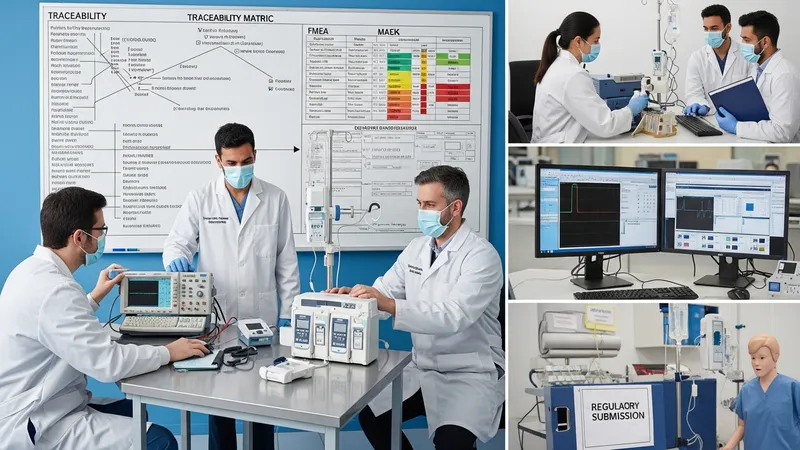
Design verification and validation commonly follow documented protocols and acceptance criteria. Verification may involve bench testing and analytical checks to confirm that design outputs meet specified inputs. Validation may include simulations or clinical-style evaluations to confirm the device performs as intended in realistic settings. Software-containing devices often require distinct verification and validation artifacts, such as unit tests, integration tests, and system-level testing. Documentation of test methods, pass/fail criteria, and results typically forms part of a design dossier submitted for regulatory review or retained for internal compliance.
Risk-management activities may rely on recognized methodologies such as failure modes and effects analysis (FMEA) or fault tree analysis to structure assessments. These techniques can help manufacturers quantify risk priority and select appropriate risk controls. Risk acceptability is often considered in context—factors such as severity of harm, probability of occurrence, and availability of alternate treatments can affect decisions. Manufacturers may periodically review risk files as part of design changes, post-market data analysis, or when new hazard information becomes available, ensuring that mitigation measures remain aligned with current knowledge.
Interdisciplinary collaboration is commonly important in design and risk work. Inputs from clinicians, human factors specialists, production engineers, and quality/regulatory staff may shape decisions about materials, labeling, or production tolerances. Early engagement with manufacturing teams can also reduce later rework by ensuring designs are suited to intended production processes and capabilities. These considerations may shorten development cycles and improve consistency, while remaining procedural and verifiable for regulatory documentation purposes.
A formal quality management system (QMS) often provides the organizational framework that supports consistent device production and regulatory compliance. A QMS may define procedures for document control, change management, nonconforming product handling, and corrective actions. Manufacturers commonly maintain device master records, design history files, and batch records that together create an auditable trail from design inputs to finished product release. External audits or assessments by notified bodies or regulatory agencies may evaluate these records to confirm consistent application of the QMS.
Document control and record retention practices are frequently emphasized within compliance regimes. Procedures typically specify how documents are approved, revised, and archived, and they define retention periods for critical records. Design files may include version-controlled drawings and specifications, while production records may capture equipment logs, inspection results, and personnel training. Consistent labeling and unique identifiers, such as lot numbers or serial numbers, commonly enable traceability across supply chains and facilitate investigations when deviations or complaints occur.
Supplier management usually forms part of the QMS scope and may include qualification criteria, periodic assessments, and documented agreements on quality expectations. For critical components or services, manufacturers often require evidence of supplier controls, such as certificates of analysis, supplier audits, or performance testing results. Where suppliers provide software, sterile processing, or testing services, clear contracts and technical specifications may be used to ensure that external activities integrate with the manufacturer’s quality processes and regulatory responsibilities.
Change control is a typical QMS element used to manage design or process modifications that may affect device safety or performance. Change control procedures often require risk assessment, verification activities, and update of relevant documentation before changes are implemented at scale. Manufacturers may review the impact of changes on regulatory submissions or conformity evidence and, where necessary, coordinate notifications with regulatory authorities according to applicable rules. These structured practices help maintain compliance over the device life cycle.
Manufacturing processes for medical devices can vary from manual assembly to highly automated lines, depending on device complexity and production volume. Process qualification activities—such as equipment installation qualification (IQ), operational qualification (OQ), and performance qualification (PQ)—may be used to document that equipment and processes perform consistently. Environmental controls for cleanrooms, laminar flow zones, or controlled storage are often specified when devices are sensitive to particulate or microbial contamination, and monitoring data may be retained as part of release documentation.
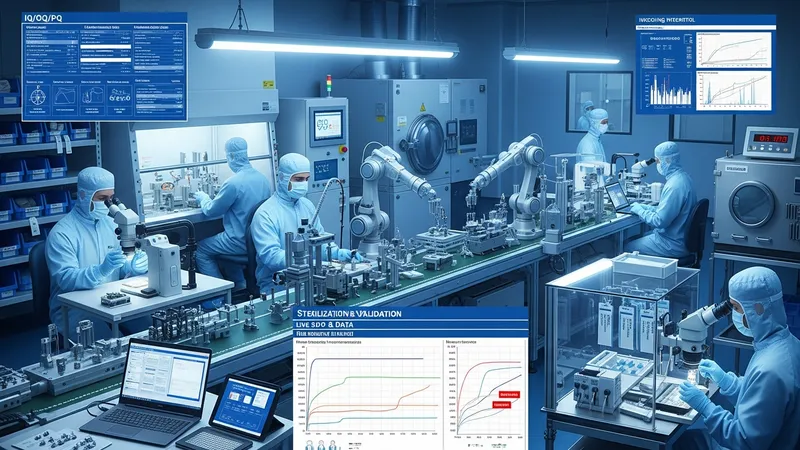
Component inspection and in-process controls typically support consistent quality. Incoming inspection criteria, in-process sampling plans, and final release tests can be defined based on device risk and criticality of components. Statistical process control methods may be used to monitor key parameters and identify trends before they lead to out-of-specification product. For devices requiring sterilization, validation protocols for sterilization cycles and sterility assurance level targets are commonly documented, and packaging integrity tests may be performed to confirm barrier performance.
Supply chain resilience and traceability are often areas of regulatory focus. Manufacturers may maintain bills of materials that list approved suppliers and part numbers, and they may require suppliers to notify them of changes that could affect performance. Where single-source components are used, contingency planning and alternative qualified suppliers are considered as risk mitigations. Batch or lot tracking systems can facilitate recalls or targeted communications if a distributed component is later found to be nonconforming.
Testing protocols are commonly tailored to the device’s intended function and applicable standards. Bench testing may cover mechanical durability, electromagnetic compatibility, or software performance, while biocompatibility assessments may be necessary for devices with tissue contact. Manufacturers often compile test reports and certificates from accredited laboratories to support technical documentation. These results may be used for internal release decisions and to satisfy external conformity assessments as required by regulators or standards bodies.
Regulatory pathways for medical devices typically depend on device classification, intended use, and associated risk. Manufacturers may follow different conformity assessment routes such as self-declaration, third-party certification, or submission of clinical evidence, depending on regional rules and device risk categories. Early determination of the applicable classification and conformity procedures can help define the scope of required documentation, testing, and potential interactions with regulatory authorities. Manufacturers may also consider international regulatory requirements when planning market access in multiple jurisdictions.
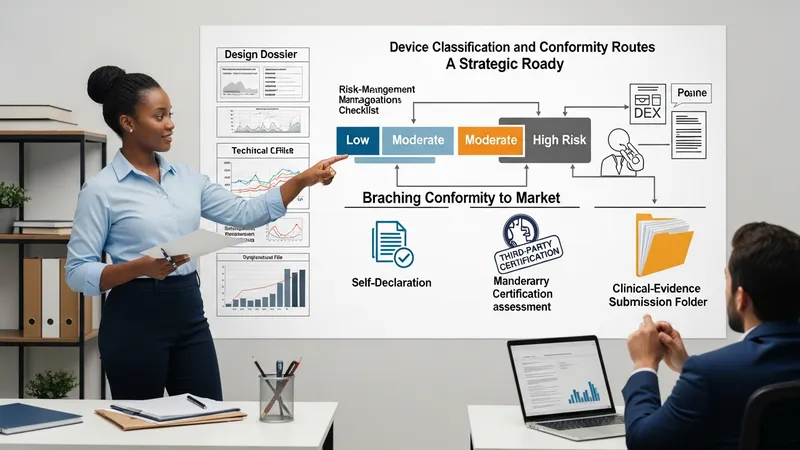
Market access planning often includes preparation of a technical file or design dossier that summarizes design rationale, verification and validation evidence, manufacturing controls, and risk management. For some devices, clinical data or literature may be part of the dossier to demonstrate performance under intended use. Manufacturers may maintain a regulatory affairs function to monitor changes in standards and regional regulations, since updates can influence labeling, post-market obligations, or required conformity assessments over time.
Post-market surveillance and vigilance processes are commonly required to continue assessing device performance after commercialization. Activities may include complaint intake procedures, adverse event reporting to competent authorities as applicable, periodic safety update reports, and trend analysis to detect changes in performance metrics. Corrective and preventive actions (CAPA) procedures provide structured methods to investigate root causes and document measures taken. These outputs can feed back into design, manufacturing, or labeling changes as part of continuous compliance management.
When devices are modified or new information emerges, manufacturers may reassess regulatory obligations and update technical documentation accordingly. Changes that affect safety or intended use may trigger additional testing, reclassification, or renewed conformity assessments. Manufacturers often document decision rationales and the associated evidence to demonstrate that regulatory requirements were considered during change implementation. Continued alignment of design, production, and post-market data supports sustained conformity across the product life cycle.